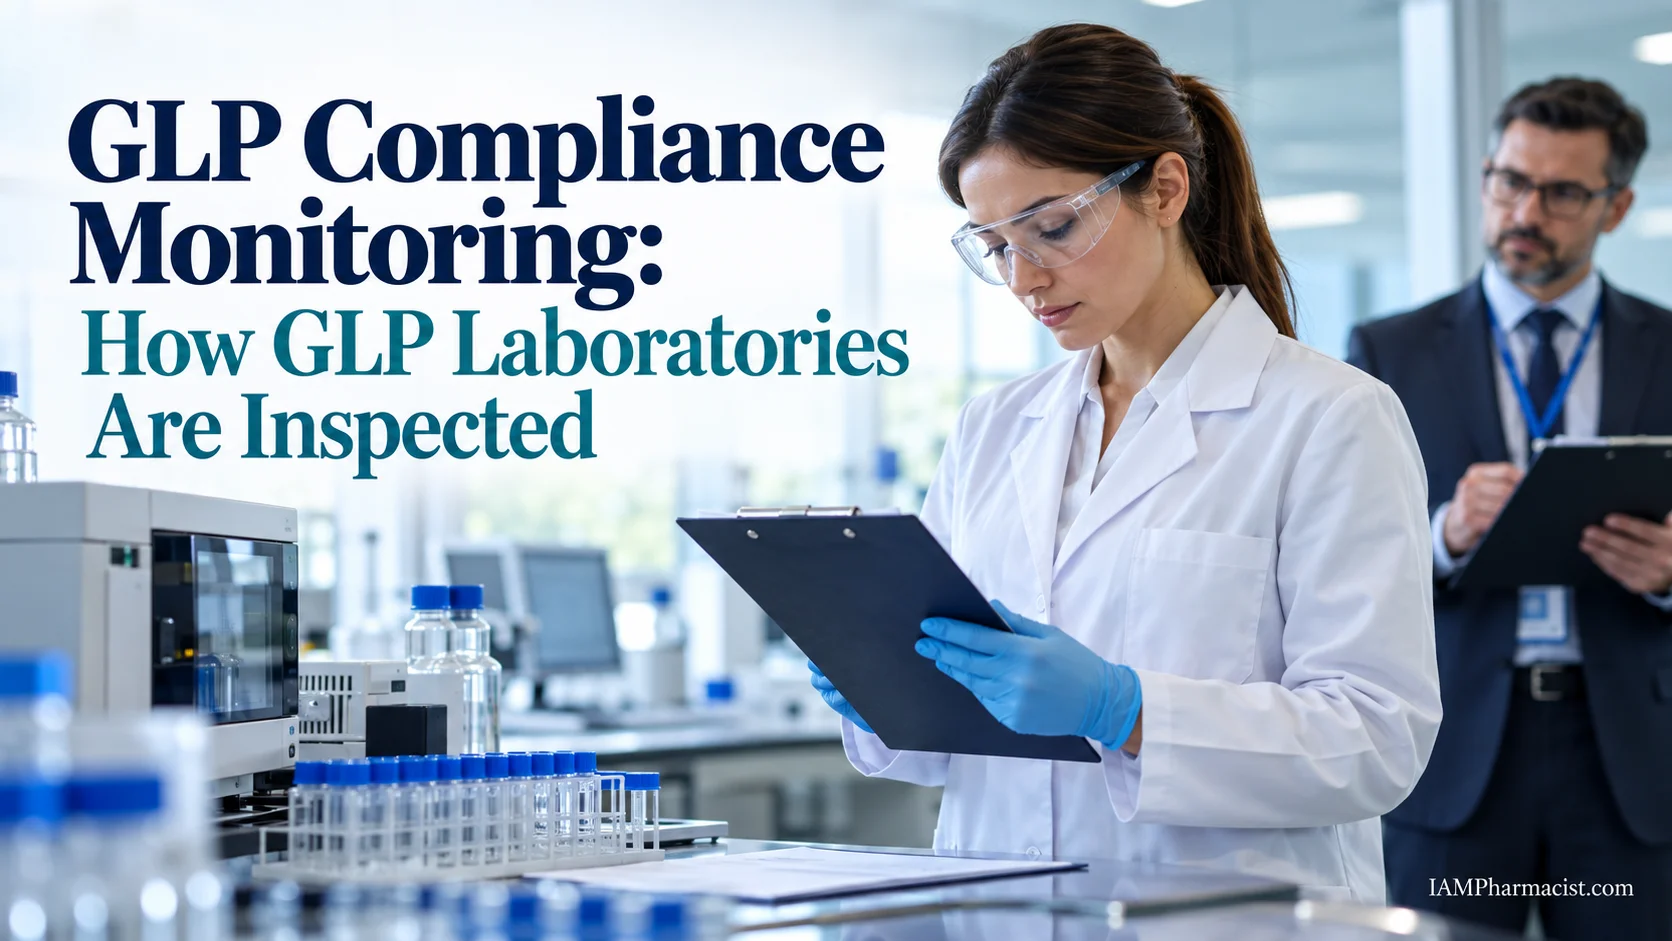

Learn how GLP Laboratories Are Inspected, what FDA, EPA and OECD inspectors review, and how laboratories can prepare reliable study records.
GLP compliance monitoring is the process authorities use to check whether nonclinical health and environmental safety studies are planned, performed, recorded, reported, and archived under the applicable Good Laboratory Practice requirements. Monitoring may include an inspection of a test facility, an audit of selected studies, or both. The exact requirements depend on the authority and the study’s regulatory purpose. The OECD describes compliance monitoring as national-program inspections and study audits; it does not create one worldwide laboratory certificate or inspection timetable.
For pharmaceutical professionals, GLP is most relevant to nonclinical studies that support regulatory submissions, such as certain toxicology studies. It is distinct from GMP, which governs the manufacture and quality control of medicines. A laboratory’s routine product-release testing is not automatically a GLP study; first determine which rules apply to that work. https://iampharmacist.com/gmp-vs-cgmp/
Which GLP Framework Applies?
The applicable framework depends on the product, study purpose, and receiving authority. The following are common reference points:
| Framework | Main application |
|---|---|
| OECD GLP Principles | Principles for organizing and conducting nonclinical safety studies. National compliance-monitoring programs inspect facilities and audit studies under their local arrangements. |
| FDA, 21 CFR Part 58 | Nonclinical laboratory studies supporting or intended to support FDA research or marketing permit applications, including studies involving drugs, biologics, and human-use medical devices. |
| U.S. EPA, 40 CFR Part 160 | Studies supporting applications for research or marketing permits for pesticide products regulated by EPA. |
| U.S. EPA, 40 CFR Part 792 | Certain health-effects, environmental-effects, and chemical-fate studies under the Toxic Substances Control Act (TSCA). |
These frameworks share a focus on study quality and data integrity, but their legal scope and detailed provisions are not interchangeable. Check the actual regulation and guidance for the study rather than relying on a generic comparison.
Types of GLP Inspection and Study Audit
A routine or surveillance inspection assesses a facility’s current GLP operations and may include selected ongoing or completed studies. FDA’s inspection program describes both surveillance and directed assignments; FDA investigators use the study schedule and inspection scope to select studies for review. The program does not establish a universal rule that every facility is inspected every one or two years.
A directed or for-cause inspection is initiated for a specific reason, such as concerns about study data or reported practices. A study audit traces a selected study through its protocol, raw data, records, and final report. It may take place as part of a broader inspection rather than as a separate inspection category.
What Happens During a GLP Inspection?
The sequence varies by authority and assignment, but an inspection commonly includes these activities:
| Stage | What may happen |
|---|---|
| Scope and opening discussion | Inspectors explain the inspection’s purpose and scope. Facility representatives identify key contacts, relevant studies, and organizational changes. |
| Facility walkthrough | Inspectors assess whether the facilities, equipment, and work areas are suitable for the studies being conducted. |
| Records and study review | Inspectors examine selected protocols, records, raw data, SOPs, training records, equipment logs, and Quality Assurance Unit (QAU) documentation. |
| Closing discussion and follow-up | Inspectors discuss preliminary observations. Any written response or corrective action follows the applicable authority’s process and deadlines. |
The FDA inspectional program describes review of facilities, study selection, protocols, study conduct, final reports, and records. Inspectors may also check that study protocol changes are signed, dated, and maintained with the protocol.
Records and Controls Inspectors Commonly Check
Inspection focus depends on the study, but laboratories should be ready to show how their records support reliable results:
- Personnel and responsibilities: Training, experience, job descriptions, and assigned study duties.
- Facilities and equipment: Suitability for the study, equipment maintenance and calibration records, and controls that prevent mix-ups or interference.
- Quality Assurance Unit: Under FDA Part 58 and EPA Part 792, the QAU must be separate from and independent of personnel directing or conducting the study. Inspectors may examine its inspection records and follow-up.
- Protocol and study conduct: Whether the study followed its approved protocol and SOPs, and whether changes or deviations were authorized and documented.
- Test and control articles: Where applicable, records establishing identity, strength, purity, stability, and uniformity, along with preparation and handling information.
- Raw data, reports, and archives: Whether the final report accurately reflects the underlying study records, and whether required material can be retrieved from its archive. FDA rules allow a testing facility to use a contracted commercial archive; archive arrangements and retention periods must follow the applicable requirements.
Practical Pharmaceutical Example
Suppose a laboratory conducts a nonclinical oral toxicity study for a drug candidate. An inspector could trace the test article from receipt and identification through formulation preparation, dosing, observations, sample analysis, and the final report. If a scheduled observation was missed, the inspector may check whether the event was recorded promptly, assessed for its effect on the study, and handled under the protocol and applicable procedures. Records should make the history clear; undocumented reconstruction weakens the evidence.
Preparing for an Inspection
Maintain an up-to-date study schedule, controlled SOPs, current training records, equipment logs, QAU inspection reports, and a clear archive index. Ensure staff know how to locate original records and who is authorized to answer questions. Before the visit, check that selected study records agree across the protocol, raw data, deviations, and final report.
If an observation requires action, document the issue, its cause and scope, the correction, and steps to reduce recurrence. Track completion and assess whether the action worked. Do not assume that every GLP authority uses the same CAPA form, response deadline, or inspection outcome. When FDA issues a Form 483, it records inspectional observations; the form itself is not a final agency determination.
Key Takeaways
- GLP compliance monitoring may involve facility inspections, study audits, or both.
- The governing requirements depend on the study and receiving authority.
- Inspectors commonly trace records from the protocol and raw data to the final report.
- There is no universal inspection interval or single GLP certificate that applies worldwide.
- Corrective actions should be documented and aligned with the applicable authority’s process.
Frequently Asked Questions
1. What does GLP compliance monitoring assess?
It checks whether relevant facilities and nonclinical studies follow the applicable GLP requirements and whether study data are reliable and traceable.
2. Which agencies inspect GLP laboratories?
Depending on the study, monitoring may be conducted by FDA, EPA, or a national GLP compliance-monitoring authority operating under its country’s framework.
3. What is the difference between routine and directed GLP inspections?
Routine or surveillance inspections assess general compliance and selected studies. Directed inspections focus on a particular concern or assignment.
4. How often do GLP inspections take place?
There is no universal one-to-two-year interval. Inspection frequency depends on the relevant authority’s program and the facility or study circumstances.
5. What records are reviewed during a GLP inspection?
Inspectors may review protocols, raw data, final reports, SOPs, training records, equipment records, QAU documentation, and archive records.
6. Must the Quality Assurance Unit be independent?
FDA Part 58 and EPA Part 792 require the QAU to be separate from and independent of personnel directing or conducting the study.
7. Is every study externally audited?
Not necessarily. Authorities select studies according to the inspection’s scope and applicable program; a laboratory’s internal QAU has separate monitoring duties.
8. Can a GLP archive be off-site or contracted?
FDA Part 58 allows a facility to contract with a commercial archive. Confirm that the specific archive arrangement meets the applicable rule and supports retrieval and retention.
9. What should a laboratory do after an inspection finding?
Assess the finding, determine its scope and cause, document appropriate corrections and preventive steps, and follow the authority’s response process and deadline.
10. Is GLP the same as GMP?
No. GLP applies to the organization and conduct of applicable nonclinical studies. GMP applies to manufacturing and related product-quality controls.
Official External References
- FDA: 21 CFR Part 58—Good Laboratory Practice for Nonclinical Laboratory Studies
- FDA: GLP Inspectional Program
- EPA: GLP Standards Compliance Monitoring Program
- EPA: 40 CFR Part 160—Good Laboratory Practice Standards
- EPA: 40 CFR Part 792—Good Laboratory Practice Standards
- OECD: Good Laboratory Practice and Compliance Monitoring