
Learn GLP archives requirements for study records, specimens, retention periods, access controls and electronic data migration.
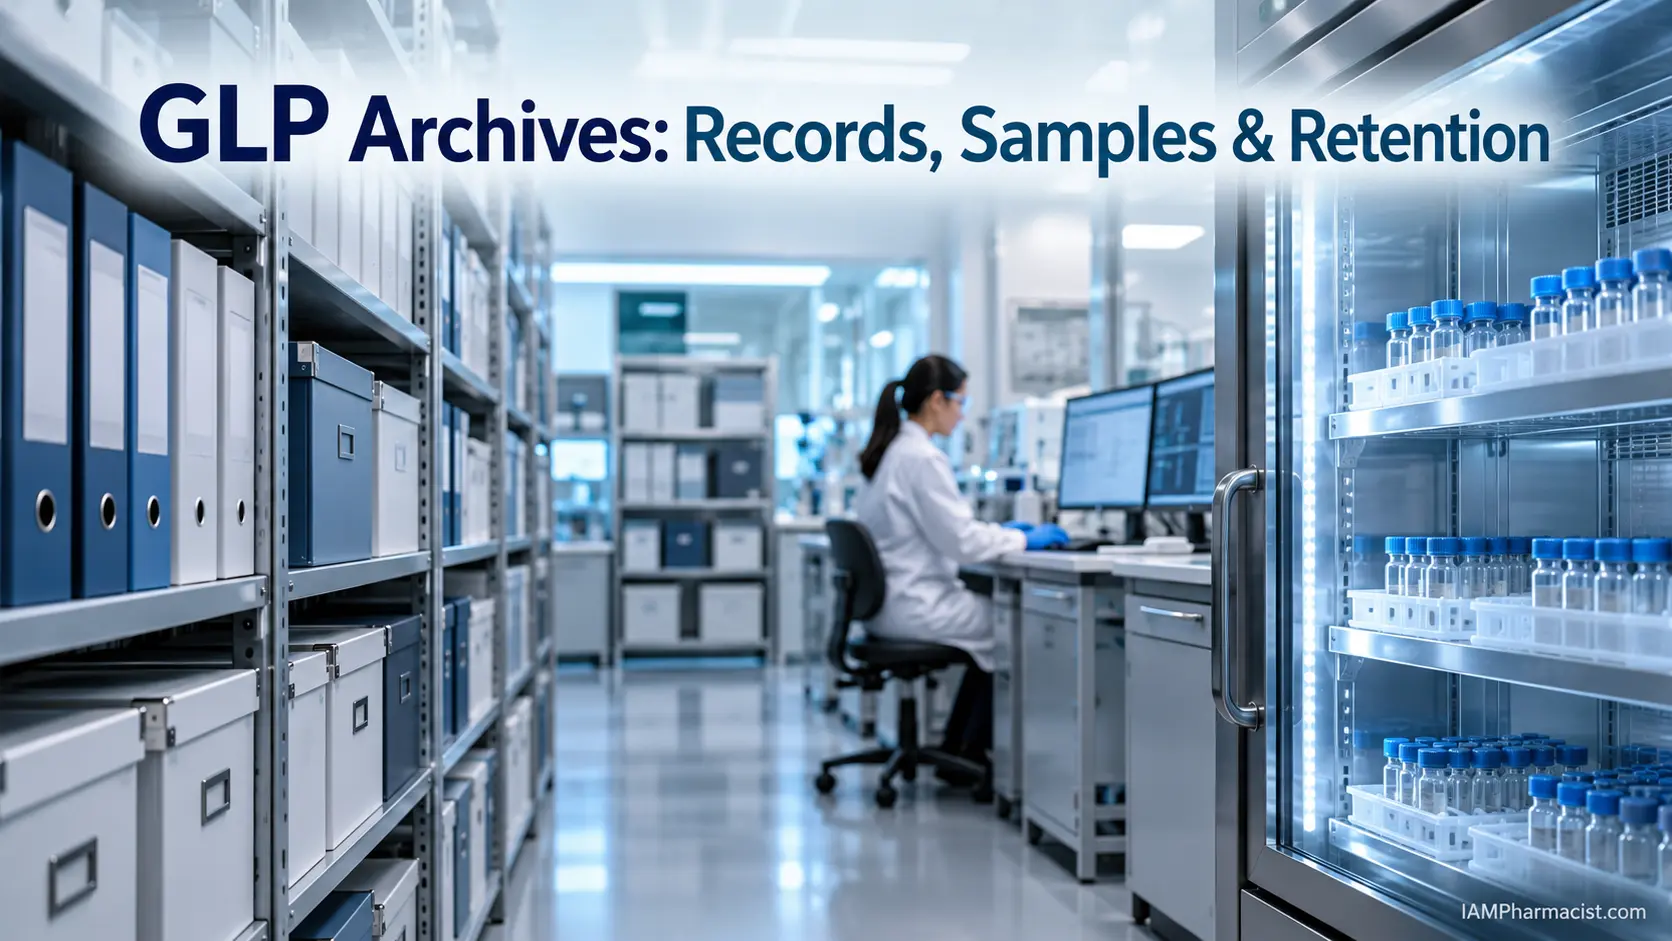
A Good Laboratory Practice (GLP) archive protects the records and materials needed to review and reconstruct a nonclinical safety study. It may contain paper records, electronic raw data, protocols, reports, and study specimens. FDA’s 21 CFR Part 58 requires orderly storage, prompt retrieval, a designated person responsible for the archives, restricted entry, and indexing. OECD guidance adds practical recommendations for managing archive access and electronic data over time.
Key takeaways
- FDA GLP retention periods depend on whether a study supports an FDA application or is completed without such a submission.
- 21 CFR Part 58 does not set a universal 10–15-year retention period.
- Certain specimen types and fragile materials have specific retention exceptions or limits.
- A named person must be responsible for the FDA archive, but the regulation does not require that person to be an independent, full-time archivist.
- Electronic migration should preserve data meaning, associated metadata, and information needed to reconstruct the study.
What should a GLP archive contain?
Under 21 CFR § 58.190, records and materials generated by a nonclinical study generally include raw data, documentation, protocols, final reports, and specimens. Raw data can be paper or electronic. For example, a pharmaceutical toxicology study may generate signed worksheets, instrument files, histology slides, dose formulation results, and a final report.
Other supporting records have specific retention provisions. The Quality Assurance Unit’s master schedule, protocol copies, and inspection records must be maintained in an accessible record system. Equipment maintenance and calibration records are retained for the applicable period. Training and experience summaries and job descriptions may be retained with employment records. Part 58 requires historical SOP files, but does not say every supporting record must be stored in the same physical archive as the study file.
Some specimens are treated differently. Section 58.190 excludes certain specimens—including those from mutagenicity tests and wet blood, urine, feces, and biological-fluid specimens—from its general retention requirement. Section 58.195 also addresses fragile specimens, test or control article samples, and specially prepared materials. Facilities should classify each material and apply the relevant rule rather than assume every sample must be kept for the same duration.
Archive access, storage, and retrieval
FDA requires a person to be identified as responsible for the archive; only authorized personnel may enter, and materials must be indexed so they can be retrieved. Storage conditions must minimize deterioration based on the material and retention period. This may mean different controls for paper reports, frozen biological samples, and glass microscope slides.
A practical archive procedure should identify who can request, authorize, remove, and return a record or specimen. Keep a movement log for materials taken out for review and returned afterward. OECD archive guidance recommends recording the movement of records and materials and using procedures that allow the archivist to track them. Assess risks such as fire, water damage, temperature excursions, and humidity according to the archive and the materials stored.
Use controlled SOPs for archive requests, access, indexing, environmental monitoring, and disposition. The goal is to make a complete study record retrievable without changing the original evidence.
FDA and OECD retention periods
The FDA rule sets specific minimum periods. For applicable records, 21 CFR § 58.195 uses the shortest of the listed periods: at least two years after FDA approves the application supported by the study, or at least five years after the study results are submitted to FDA. For other situations, records must be retained for at least two years after the study is completed, terminated, or discontinued. The rule includes an exception for studies supporting INDs and IDEs, which follow the five-year submission period in § 58.195(b)(2). Other applicable record requirements may still apply.
OECD GLP does not establish one universal 10–15-year period. Retention periods are set by applicable authorities. OECD’s archive guidance strongly recommends retaining records for at least three inspection cycles if no applicable authority has defined a period; this is guidance, not a fixed number of years.
| Topic | FDA GLP: 21 CFR Part 58 | OECD GLP guidance |
|---|---|---|
| Retention period | Periods in § 58.195, subject to other applicable requirements | Set by the relevant authority; at least three inspection cycles is recommended if no period is defined |
| Archive responsibility | A person must be identified as responsible | Guidance describes archive management and retrieval controls |
| Electronic records | Part 11 controls apply when the electronic record or signature is within Part 11’s scope | OECD guidance addresses computerized systems, data integrity, and archives |
| Specimens | General rule has specified exceptions and limits for some materials | Apply applicable national requirements and archive guidance |
Electronic GLP data and migration
Electronic archives must preserve more than a readable printout when the study depends on native data, metadata, processing history, or audit trails. If electronic records fall within 21 CFR Part 11, its controls include secure, time-stamped audit trails retained at least as long as the associated records. OECD’s GLP Data Integrity guidance recommends that data migration be justified, designed, and validated to preserve meaning and integrity, including associated metadata.
Before a migration, document the source and destination, data formats, metadata, mapping rules, transfer method, and checks used to compare source and destination records. Preserve links such as the relationship between an analytical result, its operator, timestamp, instrument, and audit trail. Record the migration date and any exceptions. Keep the source records under control until reconciliation and approval are complete; define a recovery approach in case the transfer fails.
A non-proprietary format such as PDF/A or XML can help with long-term readability when suitable, but converting native analytical data to a static file may remove information needed to review or reconstruct the result. Choose the format based on the record’s use and retain the software or supporting information needed to interpret the data. Periodic retrieval and readability checks are sensible controls; set their frequency through procedures and risk assessment. OECD guidance treats migration as a data-integrity activity across the record lifecycle.
GLP archive audit checklist
- Is a person formally identified as responsible for the archive?
- Are access limited to authorized personnel and records indexed for retrieval?
- Can staff locate the study protocol, raw data, final report, and applicable specimens?
- Are removal, return, and transfer of archived materials documented?
- Are retention periods checked against the applicable country and product requirements?
- Do migrated electronic records retain relevant metadata and audit-trail information?
- Can staff retrieve and interpret archived files using available systems?
Frequently asked questions
1. What is a GLP archive?
It is the controlled location or system for storing and retrieving nonclinical study records and materials required for later review and reconstruction.
2. What records must be archived under FDA GLP?
Generally, raw data, documentation, protocols, final reports, and specimens generated by the study, subject to the exceptions in 21 CFR § 58.190.
3. How long must FDA GLP records be retained?
For applicable records, § 58.195 sets periods based on FDA application approval, submission of study results, or study completion, termination, or discontinuation. Apply the shortest listed period and check for other applicable requirements.
4. Does OECD require 10–15 years of GLP retention?
No single 10–15-year period applies across OECD GLP. The relevant authority sets retention periods; OECD guidance recommends at least three inspection cycles if no period is defined.
5. Must every biological sample be retained?
No. FDA Part 58 includes specific exceptions and limits for certain wet specimens and fragile materials. Determine the rule for each specimen type.
6. Does FDA require an independent archivist?
Section 58.190 requires an identified person responsible for the archive. It does not require that person to be an independent, full-time archivist.
7. Should archive access and material movement be logged?
FDA requires authorized archive access and indexed records. OECD guidance recommends documenting movement of materials into and out of the archive; a movement log is a practical way to do this.
8. Are electronic records subject to 21 CFR Part 11?
Part 11 applies when electronic records or signatures are within its scope, including records maintained electronically to meet FDA requirements. It does not replace the underlying GLP record requirements.
9. Can GLP raw data be migrated to a new system?
Yes, if the migration preserves the data’s meaning, integrity, and relevant metadata. OECD guidance recommends a justified, designed, and validated migration process.
10. Is PDF/A enough to archive laboratory data?
Not always. A static PDF may be useful for viewing but may not preserve native data, metadata, or processing history needed to review or reconstruct an analytical result.
Relevant internal links
- Link ALCOA+ to the data integrity principles article.
- Link SOPs to the pharmaceutical SOP guide.